1 Laboratory of Fundamental and Applied Physics (LPFA), University of Abobo Adjamé, (Now Nangui Abrogoua), Abidjan, Côte d’Ivoire;
2 Laboratory of Structural and Theoretical Organic Chemistry, University of Félix Houphouët-Boigny, Abidjan, Côte d’Ivoire;
3 International Center for Theoretical Physics, ICTP-UNESCO, Coastal Road, Trieste, Italy.
4 International center for applied research and sustainable technology, Bratislava, Slovaquie.
5 Laboratory of Crystallography-Molecular Physics, University of Félix Houphouët-Boigny, Abidjan, Côte d’Ivoire.
* Corresponding Author
ORCID Details
Kouassi F: 0009-0008-7629-8575
Kouakou J-L.: 0009-0006-3725-433X
Ané M.: 0009-0003-2665-7414
Kouman C: 0009-0003-8591-4804
Kéita M: 0009-0004-5113-9712
Megnassan E: 0000-0003-1505-5277

World Journal of Advanced Research and Reviews, 2026, 31(03), 626–640
Article DOI: 10.30574/wjarr.2026.31.3.2344
Received on 02 August 2026; revised on 05 September 2026; accepted on 07 September 2026
Lysine-specific demethylase 1 (LSD1) is a histone-modifying enzyme, validated as a promising anti-cancer drug development target. Substantial LSD1 inhibitors design research has been reported so far. In this work, by a rational computer-aided molecular design (CAMD) approach, a series of 23 pyrrole derivatives, LSD1 inhibitors, were docked into LSD1 active site and their affinity with the enzyme target assessed. Explains through the existing interactions between pyrrole inhibitors and LSD1 the variation of LSD1 inhibition potency substantially. The linear correlation between the experimental LSD1 inhibition biological activities( ) and the calculated binding affinity
 ) (
 0.1626
 + 8.1632, R2 = 0.96) explains 96% of the experimental LSD1 inhibition potency by that of the computed binding affinity. In the same way the calculated interaction energy between LSD1 and TPx, is correlated with
 with R2 = 0.92. The Van der Waals (VDW) component of Eint is in linear correlation with
 with a coefficient of determination R_Vdw^2=0,87 while it dropped to R2 = 0.42 for the electrostatic component. Moreover, the analysis of individual contribution of the residues within the active site to the interaction energy could provide the structural information necessary to explain the inhibitory activity drop across the three classes of ligands, namely the most active, moderately active and less active ones. Then, these pyrrole derivatives with known
 have been selected to build a pharmacophore (PH4) model exhibiting a linear correlation between (
 ) estimated by PH4 and the experimental inhibitory potential (
 ). The equation of this correlation between pIC_50^exp and PH4-estimated
 is
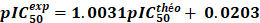 with a coefficient of determination R2=0.92. The generated 3D QSAR PH4 model is then robust for identifying new active molecules.
LSD1, QSAR, PH4, Pyrrole Derivatives, Molecular Docking, Pharmacophore, Interaction Energy.
Preview Article PDF
Affiba Florance Kouassi, Affian Mathieu Ané, Kouakou Jean-Louis KOUAKOU, Koffi Charles Kouman, Adama Niaré, Mélalie Kéïta and Eugène Megnassan. DEVELOPMENT OF AN RQSA MODEL FOR THE STUDY OF INTERACTIONS BETWEEN THIENO INHIBITORS [3, 2−B] PYRROLE DERIVATIVES (TPX) AND LYSINE-SPECIFIC DEMETHYLASE 1 (LSD1). World Journal of Advanced Research and Reviews, 2026, 31(03), 626–640. Article DOI: https://doi.org/10.30574/wjarr.2026.31.3.2344